



PRM proteomics data analysis converts scheduled high-resolution MS2 files into a peptide or protein matrix for a closed target list. Core analysis tasks are import raw data with run metadata, load the inclusion list and expected fragments, extract fragment chromatograms (XICs), review peaks and spectral consistency, choose quantifier ions, normalize across runs with QC pools or internal standards when planned, then export tables for predefined contrasts only (Peterson et al. 2012; MacLean et al. 2010). Tools such as Skyline visualize these steps; commercial suites add automation, but requirements stay tool-agnostic. Analysis does not replace study design, quantify off-panel peptides, or convert relative areas to concentration without SIS calibration. Lock software version and integration rules before group statistics.
Researchers who already have PRM raw files, a closed peptide panel, and defined groups can also review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and Targeted Proteomics Service for project-specific analysis planning.
What PRM Data Analysis Must Deliver
PRM records full MS2 for scheduled precursors. Analysis must tie each integrated trace back to the acquisition inclusion list and a declared relative or absolute claim.
Analysis task |
Minimum output |
Tool-independent QC |
|---|---|---|
Target mapping |
Precursor charge and modification state |
Panel version frozen |
Trace extraction |
Fragment XICs per peptide |
Mass accuracy windows documented |
Identity check |
Qualifier ions or spectral match |
Interference flagged |
Quantification |
Peptide-level areas or SIS ratios |
Integration rules versioned |
Normalization |
Comparable matrix across runs |
QC pool or bridge design |
Export |
Sample-by-peptide table |
Contrast arms predefined |
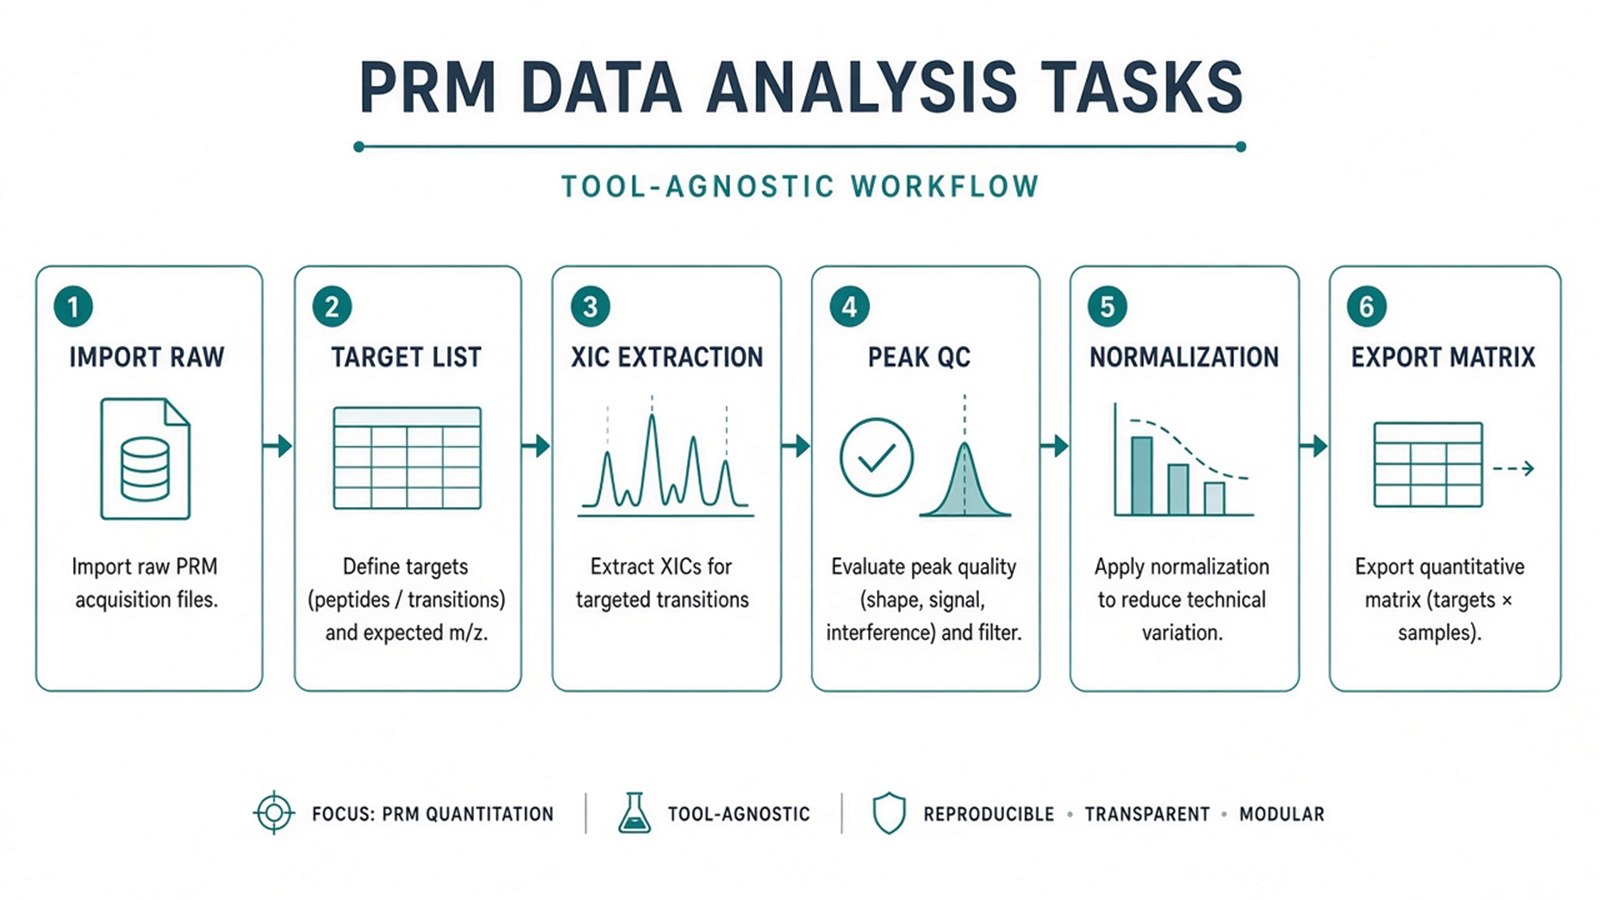
Figure 1. PRM data analysis tools differ in interface; they should all support traceable closed-panel quantification.
Choosing software before confirming PRM raw files and a frozen target list reverses the analysis order.
For step-by-step processing, see PRM Proteomics Data Processing Workflow. For mechanism context, see Mechanism of MRM/PRM in Protein Quantification.
Common Tool Classes and When Each Fits
Tools should be matched to file type, panel size, and how much manual review the matrix requires.
Tool class |
Typical strengths |
Typical limits |
|---|---|---|
Open targeted environments (for example Skyline) |
XIC visualization, manual review, multi-vendor import, library-assisted targets |
Throughput depends on operator QC discipline |
Commercial targeted suites (for example SpectroDive-class) |
Automated integration rules, report templates, batch QC |
License cost; rules still need matrix validation |
Vendor instrument software |
Tight raw-file compatibility |
May be less flexible for cross-vendor studies |
Scripted pipelines (R, Python on exported tables) |
Reproducible stats after matrix QC |
Does not replace peak review at extraction |
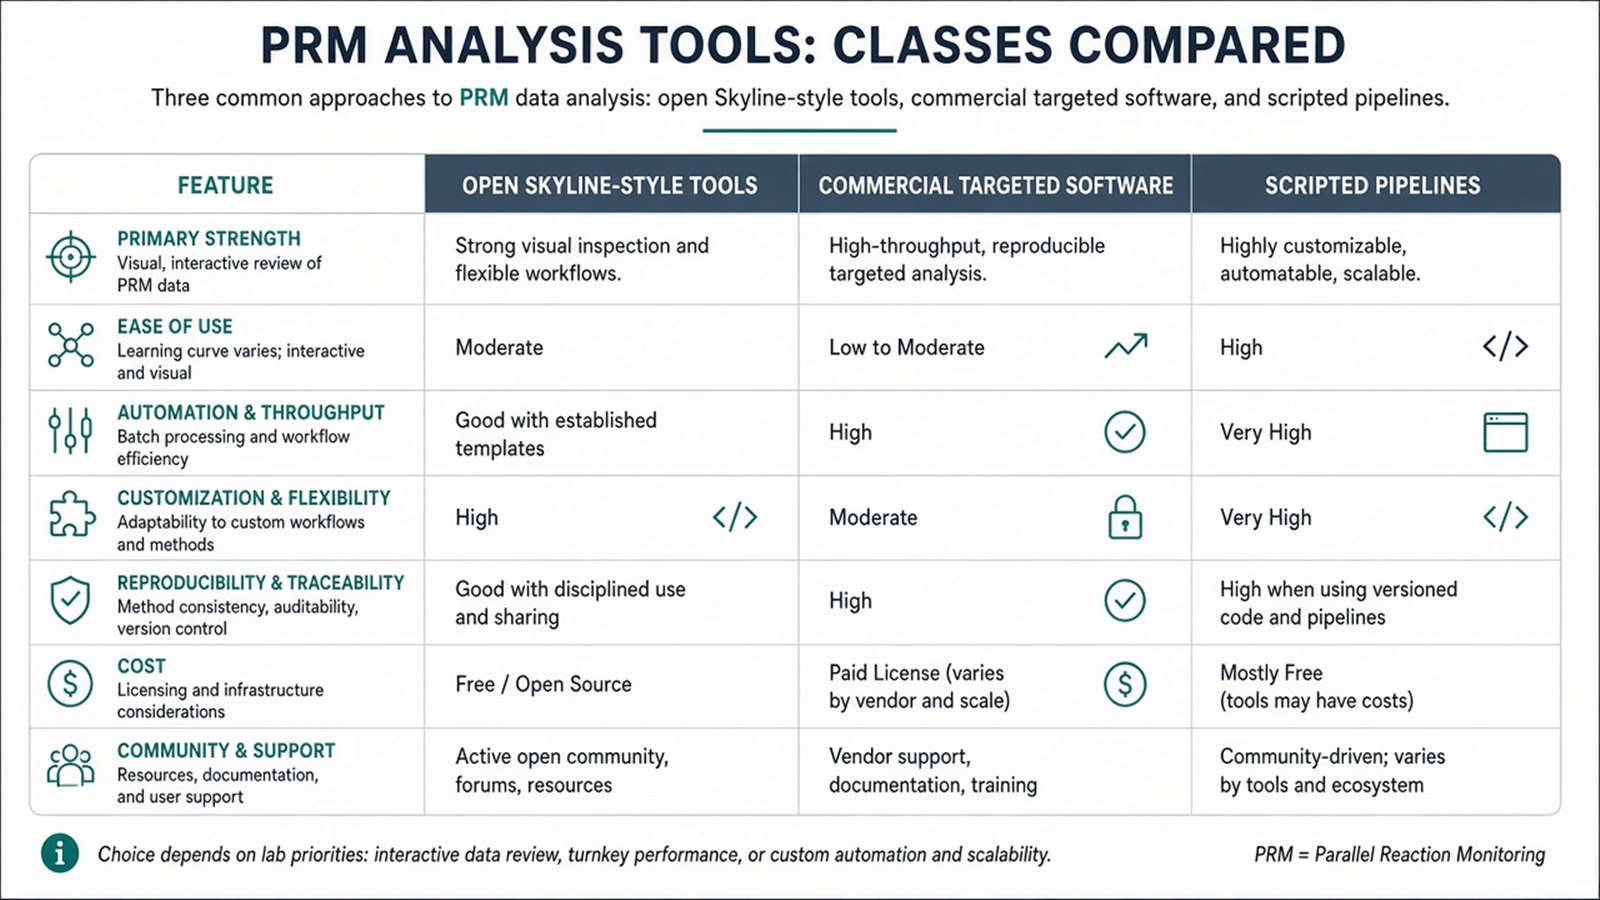
Figure 2. Tool choice follows panel closure, need for visual QC, and automation level; PRM files are not DIA discovery files.
Skyline is widely used for PRM because it supports targeted list management, chromatogram inspection, and export to external statistics (MacLean et al. 2010). SpectroDive-class tools suit standardized multi-batch verification when integration rules are validated on QC material. OpenSWATH-style pipelines primarily serve windowed DIA; use them for PRM only when the project plan explicitly maps PRM exports into a compatible targeted extraction workflow, not as a default substitute for PRM chromatogram review.
For PRM versus MRM software context, see PRM vs MRM: Comparative Analysis in Quantitative Proteomics. For discovery handoff, see Application of DIA-PRM Proteomics. For stats after export, see How to Analyze Quantitative Proteomics Data.
Judgment Criteria: Tool and Route Selection
Project state |
PRM analysis tooling often fits |
Reassess |
|---|---|---|
Files |
Scheduled HR MS2 PRM acquisitions |
DDA discovery or DIA windows only |
Panel |
Closed peptides with scheduled RT |
List still changing weekly |
Review need |
Complex matrix; visual peak QC |
Expect blind automation without pilot QC |
Absolute claim |
SIS peptides and curve logic in tool path |
Relative areas interpreted as concentration |
Scale |
Moderate panels with documented rules |
Very large multiplex better on mature MRM |
Prefer Skyline-class open tools when fragment review and transparent integration rules matter. Prefer commercial targeted suites when validated automation and reporting speed dominate. Prefer MRM vendor tools when transitions are mature on a triple quadrupole. Prefer DIA extractors only when raw data are windowed discovery files, not PRM schedules.
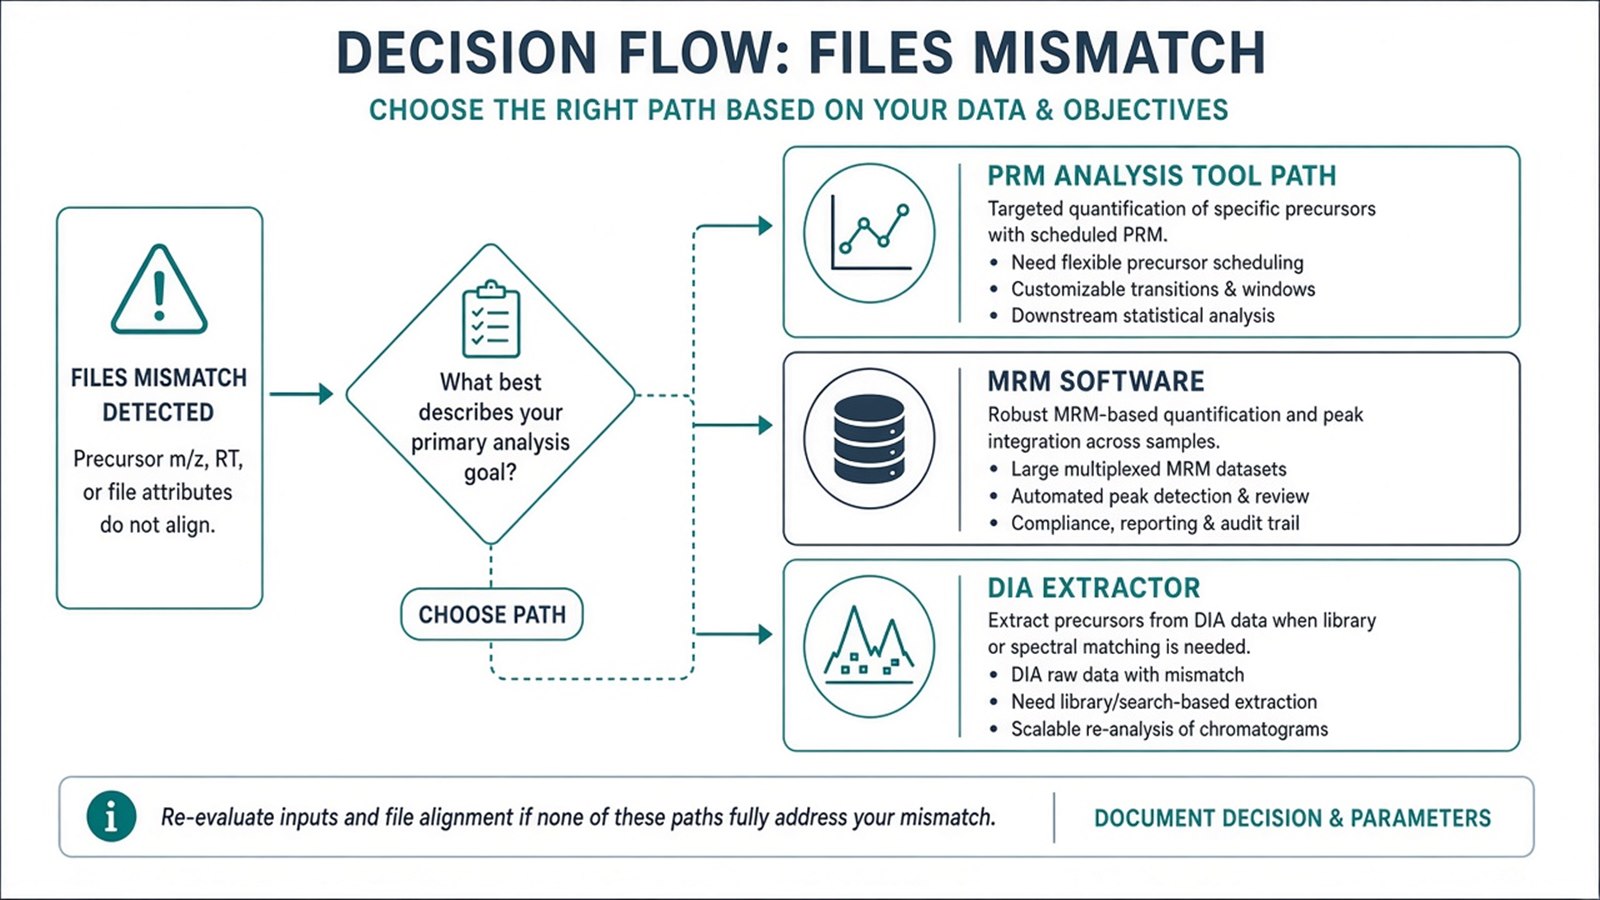
Figure 3. Match analysis tools to PRM file type and closed-panel QC needs; do not run discovery pipelines on targeted schedules by default.
For advantages at method level, see Advantages and Disadvantages of Parallel Reaction Monitoring. For validation planning, see How to Plan a Targeted Proteomics Validation Study. For absolute unit design, see Absolute Quantification in Targeted Proteomics.
Limits and Application Boundaries
Tools cannot quantify peptides that were never scheduled. Automation does not remove co-isolation interference without spectral and chromatographic QC. Relative export tables are not diagnostic assays. Software upgrades should be reported as new processing versions. Typical applications after sound PRM analysis:
- Biomarker-candidate confirmation matrices from discovery shortlists.
- Pharmacodynamic panels on fixed peptides across treatment arms.
- Biopharmaceutical monitoring with documented integration rules.
- SIS-backed amount readouts when calibration QC is included in the same pipeline.
Diagnostic cutoffs sit outside ordinary research PRM reporting unless separately validated.
Once tool class, version, and integration rules are locked, the project can deliver a traceable PRM matrix. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service for raw-file intake and analysis planning.
Frequently Asked Questions
1. Is Skyline required for PRM data analysis?
No. Skyline is a common open option, but any workflow that performs traceable XIC extraction and documented QC can fit PRM.
2. Can DIA software replace PRM analysis tools?
Not by default. DIA extractors expect windowed discovery files. PRM needs targeted schedules and closed lists.
3. What should be locked before comparing groups?
Target list version, software build, integration boundaries, normalization method, and contrast definitions.
4. When is commercial targeted software worth it?
When integration rules are validated on QC material and many batches need consistent automated reporting.
5. Where do R or Python fit?
After a reviewed peptide matrix is exported; they handle contrast testing, not substitute for peak QC at extraction.
Reference
- B. MacLean, D.M. Tomazela, N. Shulman, et al. (2010). Skyline: an open source document editor for creating and analyzing targeted proteomics experiments. Bioinformatics, 26, 966-968. https://doi.org/10.1093/bioinformatics/btq054
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
