



Planning an affinity purification–mass spectrometry (AP-MS) project means locking four decisions before the first liquid chromatography–tandem mass spectrometry (LC-MS/MS) run: bait design, controls, sample preparation, and a validation rule. Those choices decide whether the file can support a ranked candidate list, a matched mutant contrast, or a condition contrast. They do not decide whether the file will report an interactome.
Projects fail before acquisition when a tagged bait is expressed far above the endogenous level, when empty-tag arms are omitted, when lysis is copied from an unrelated recipe, or when follow-up is postponed until a long identification table appears. This page is for a tagged-bait plan. Endogenous antibody capture is a method switch, not a later amendment to the same construct sheet. For the detection chain those four decisions will later write, How AP-MS Works provides a complementary mechanism page.
Researchers who already have a tagged line and need a ranked co-enrichment table can also review the MtoZ Biolabs Affinity Purification-Mass Spectrometry Service for project-specific feasibility and analysis planning.
Write One Sentence Before the Plasmid
The objective names the bait, the system, the contrast, and the action that the shortlist is allowed to trigger. Identifying candidate partners of a tagged protein after a written stimulus is one sentence. Comparing a wild-type bait with a binding-deficient mutant under the same tag is another. Mapping everything that binds the bait is not an objective. It produces unfocused controls.
If those four elements are missing, do not order the construct or reserve the instrument. For what the finished table is allowed to claim, Understanding AP-MS: A Comprehensive Guide provides a complementary evidence page.
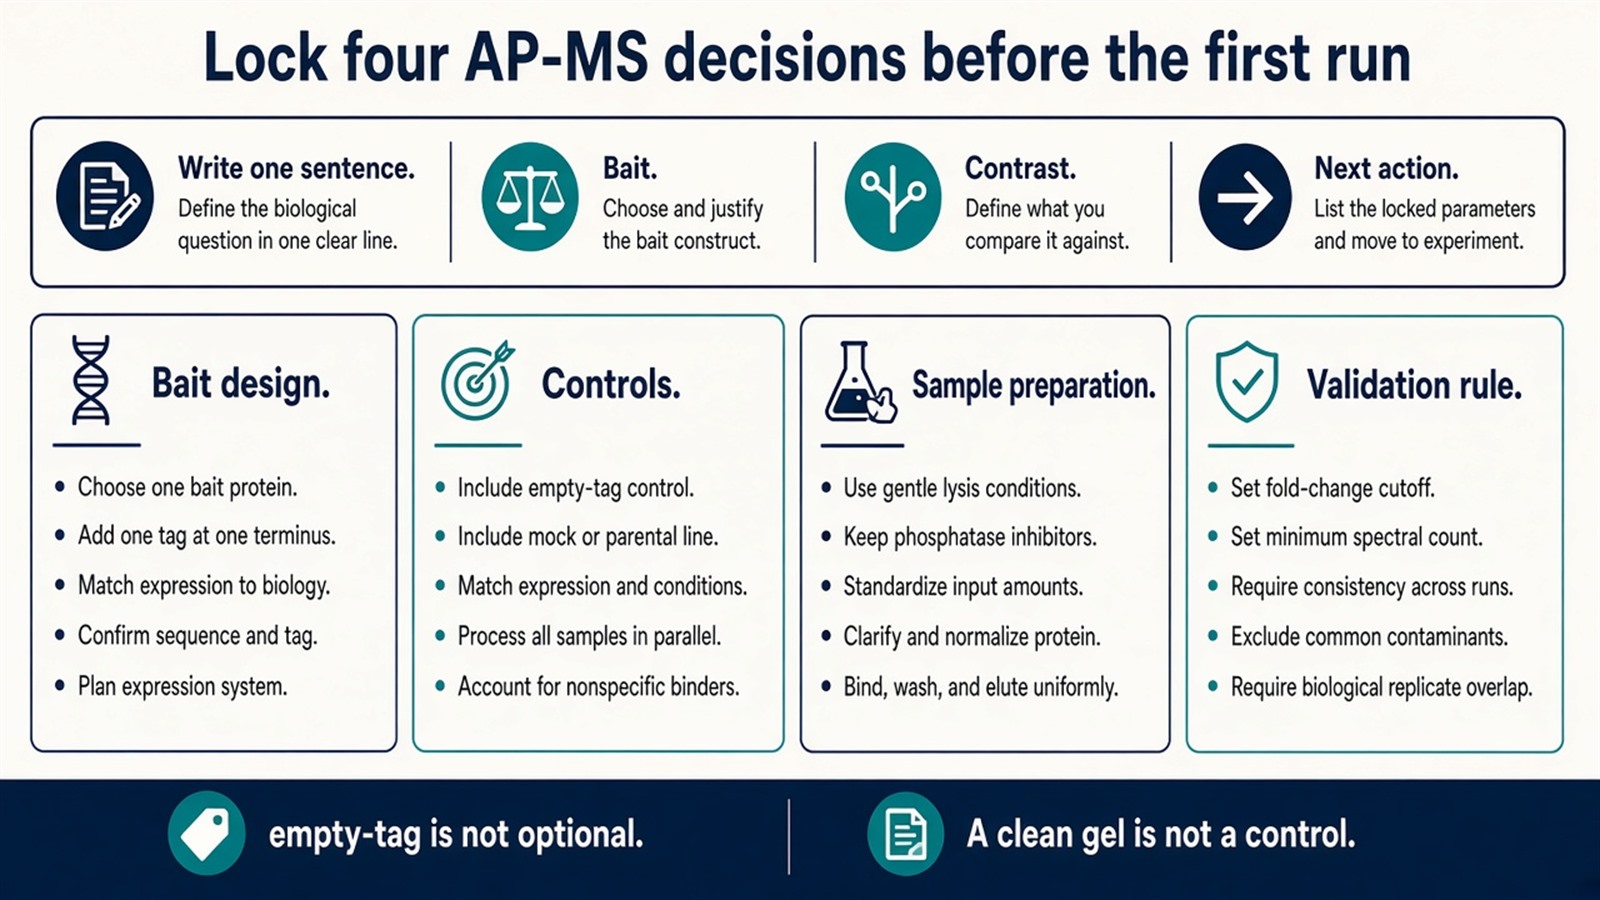
Figure 1. One sentence first: bait, contrast, next action. Then four locks. Empty-tag is not optional. A clean gel is not a control.
Write the decision the file must support before choosing FLAG or HA. A later filter cannot rebuild an arm that was not processed.
Bait Design
Common handles remain FLAG, HA, His, Strep, and GFP-class tags. Choose the chemistry that the host and the resin already tolerate. N- and C-terminal fusions can change localization, folding, or access. Position is justified by prior evidence when it exists. It is not a default that can be optimized after a failed list.
Transient, stable, or inducible expression is a dose decision. Recovery needs enough bait. Overexpression makes nonspecific names reproducible. Inducible systems lower that risk. They do not make the tagged protein endogenous.
Confirm localization or a named activity on the tagged bait before the main cohort. A mislocalized construct can enrich the same names in every replicate and still answer the wrong compartment.
Mutants, truncations, or domain swaps belong on the same tag, the same promoter, and the same host. Matched construct chemistry is an AP-MS advantage when the contrast is the mutation. It is not a general advantage over antibody IP-MS.
Controls Are Generated, Not Applied Later
Empty-tag lines mark handle and resin binders. They use the same lysis, wash, and MS depth as the bait. A clean bait gel does not replace that arm.
Untagged parental cells separate host-cell background from tag-associated background. They are useful when the expression system itself is sticky. They are not a substitute for empty-tag.
Every arm shares bead type, wash volume, time, and elution. Procedural asymmetry invents enrichment. Biological replicates stabilize ranking. A single unreplicated capture can be exploratory. It is weak support for a go-forward interaction sentence.
Binding-deficient or localization mutants can show which names depend on a surface. They do not convert remaining names into functional partners. For the broader control menu, What Controls Are Needed for Co-IP-MS and In-Cell Crosslinking MS? provides a complementary control page.
Sample Preparation Is the Wash the Claim Can Tolerate
Detergent, salt, and inhibitors are chosen for bait recovery that still leaves a filterable background. Membrane, nuclear, and labile signaling samples often need different extraction. The goal is not the cleanest gel and not partner preservation as a native inventory.
Mild washes keep weak associations and raise background. Harsh washes drop both contaminants and some true contacts. Set that stringency before the cohort so bait and control remain comparable. Plan input for the full matrix, not one tube. Incomplete clarification raises debris. Treatment windows, if any, are locked with harvest time. A condition contrast added after the chemistry is fixed is a different experiment.
Planning item |
Decision to lock |
Why it matters |
|---|---|---|
Tag and terminus |
Which tag, N or C |
Localization and access |
Expression mode |
Transient, stable, or inducible |
Overexpression artifacts |
Empty-tag line |
Yes, matched system |
Handle binders versus bait |
Replicates |
Number and biological source |
Ranking stability |
Lysis and wash |
Buffer family and stringency |
What survives to MS |
Shortlist rule |
Enrichment and replicate, not annotation |
Stops unprioritized tables |
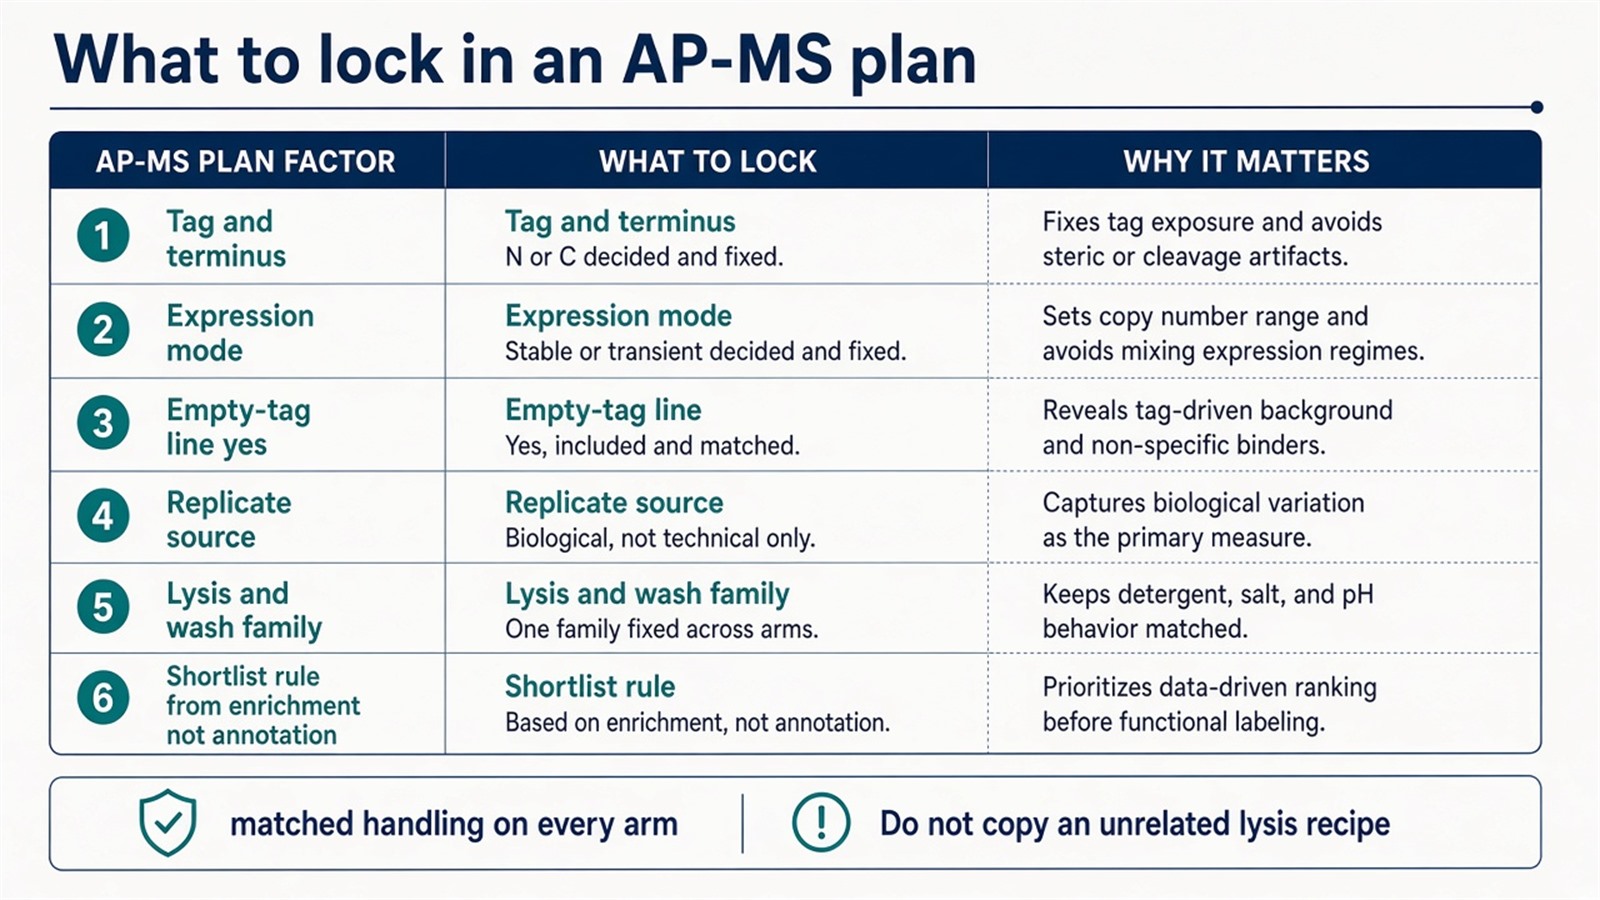
Figure 2. Six fields to write down. Matched handling on every arm. Do not copy an unrelated lysis recipe.
Validation Rules Before Acquisition
A priority candidate is defined as enrichment over empty-tag across replicates, optionally recurrent across a matched mutant or condition. Coherence with bait biology is annotation and comes last. For that read order, How to Analyze IP Mass Spectrometry Results? provides a complementary analysis-order page.
Reciprocal capture, a targeted blot, proximity labeling, or a recombinant pull-down answers different later sentences. Direct binding is not co-enrichment. A mutant-phenotype objective follows names that differ between matched baits. Weakly enriched proteins and known contaminants are parked on purpose.
Reconsider this tagged plan if the tag is incompatible, if endogenous context is mandatory and an IP-grade antibody exists, or if the real need is a recombinant pair. Those are IP-MS, Co-IP, or pull-down-MS as the first route. Switching before the cohort is a planning success.
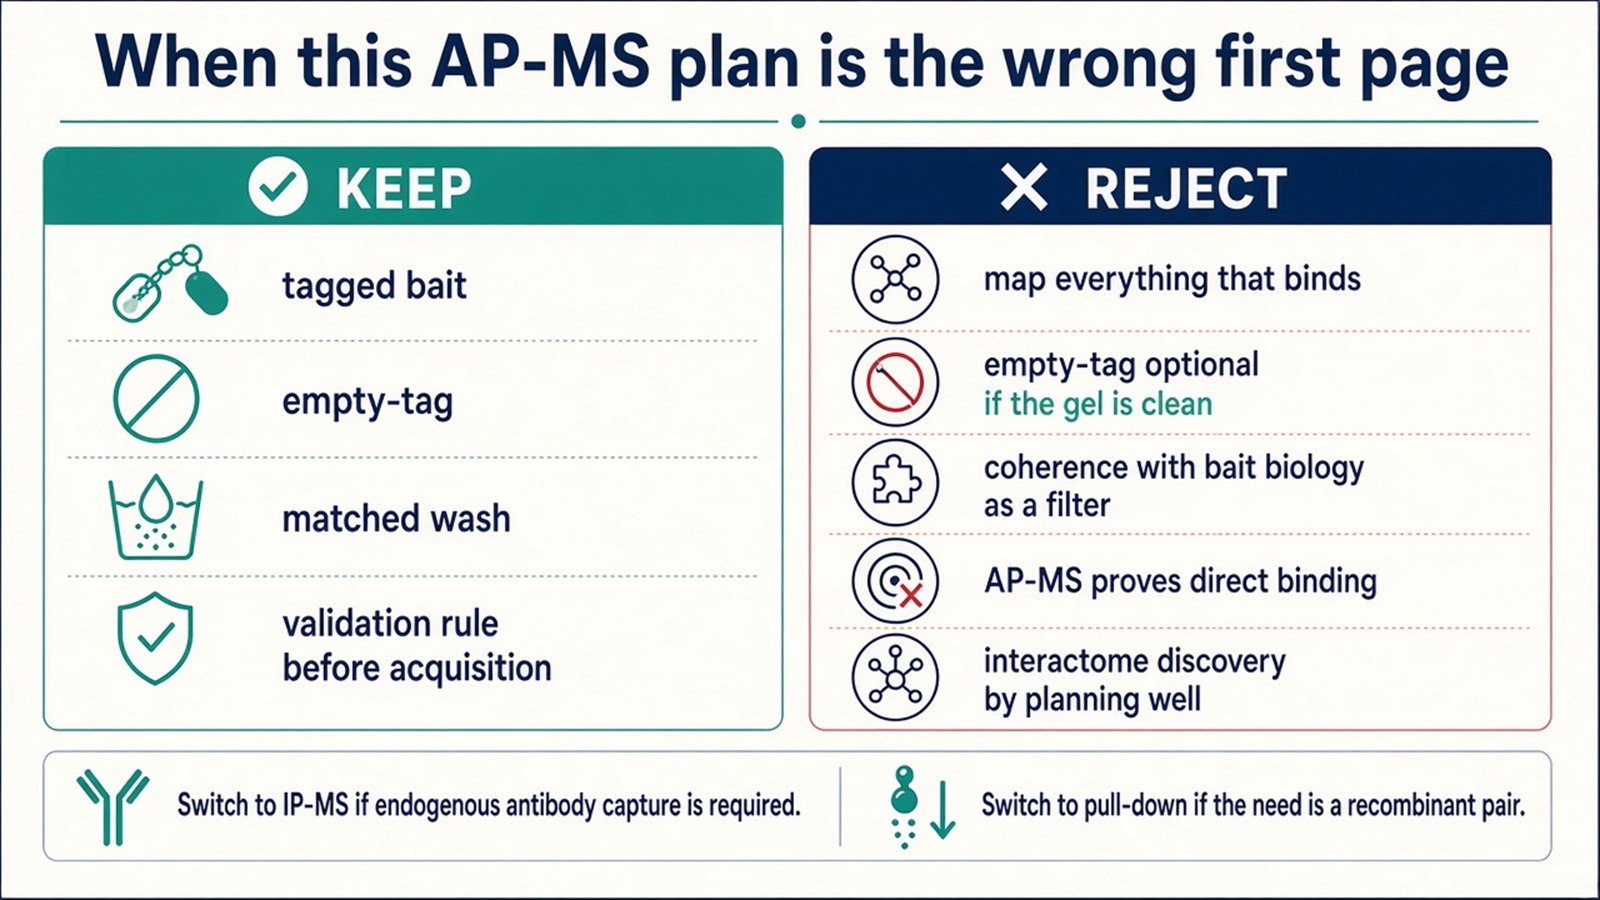
Figure 3. Keep: one sentence, empty-tag, matched wash, shortlist rule. Not granted: map everything, gel-clean as control, or AP-MS as direct-binding proof.
Experimental Limits
The plan reduces a project to one tagged bait, one control class, one wash, and one follow-up rule. Diagnostic interactome language sits outside ordinary research reporting unless a separately validated assay is already named.
Project Planning and Service Scope
A usable request names the one sentence, the tag and terminus, the empty-tag line, the replicate source, the wash family, and the orthogonal assay that would be required for the next sentence. MtoZ Biolabs scopes tagged AP-MS on Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not a planning-converts-MS-into-biology slogan.
Once the next sentence requires endogenous antibody capture rather than a tag, this plan should be closed. Researchers who need that pellet can review the MtoZ Biolabs IP-Mass Spectrometry Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What should be decided first?
The one sentence: bait, contrast, and the action the shortlist is allowed to trigger.
2. Are empty-tag controls optional if the bait gel looks clean?
No. A clean gel does not name tag- or resin-associated proteins.
3. How early should validation methods be chosen?
Before the main cohort. The rule is written with the objective.
4. Can one AP-MS plan prove direct binding?
No. AP-MS ranks co-enrichment. Direct binding is a later tier.
5. When should the plan switch to IP-MS?
When endogenous context is required and an IP-grade antibody exists.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Dunham W.H., Mullin M., Gingras A.C. (2012). Affinity-purification coupled to mass spectrometry: basic principles and strategies. Proteomics, 12, 1576-1590. https://doi.org/10.1002/pmic.201100523
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- Choi H., Larsen B., Lin Z.Y., et al. (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- Keilhauer E.C., Hein M.Y., Mann M. (2015). Accurate protein complex retrieval by affinity enrichment mass spectrometry (AE-MS) rather than affinity purification. Mol. Cell. Proteomics, 14, 120-135. https://doi.org/10.1074/mcp.M114.041012
