



Targeted protein quantification through parallel reaction monitoring (PRM) reports predefined peptides as surrogates of named proteins or modification sites. In project language, PRM on liquid chromatography-tandem mass spectrometry (LC-MS/MS) isolates scheduled precursors, records high-resolution product-ion spectra, and integrates selected fragment chromatograms on a closed panel (Peterson et al. 2012). Peptide-level readout is the usual bottom-up unit. It is not an intact-protein concentration, not open discovery label-free quantification (LFQ), and not a diagnostic cutoff by default. The guide below integrates PRM peptide quantification with workflow, locked parameters, limits, and project fit. It is not an instrument catalog, not a precision-medicine promise page, and not a software tutorial for every vendor node.
Relative change of the same peptide across predefined groups is the common follow-up claim after discovery. Amount on named peptides needs stable isotope-labeled internal standard (SIS) peptides and a calibration model in the study matrix (Gerber et al. 2003). Protein inference still depends on whether the peptide is unique and whether digest and extraction match the claim sentence (Bantscheff et al. 2012).
In MtoZ Biolabs targeted offerings, peptide-level PRM is scoped when the panel is closed, pretreatment is matched, and scheduling fits the cohort block. Service intake appears in Parallel Reaction Monitoring (PRM) Service.
What Peptide-Level PRM Covers in Targeted Quantification
Four scopes belong in the brief and stay separated.
Assay scope is targeted. Only peptides on the inclusion list are scheduled. Open search breadth is outside this record.
Peptide-surrogate scope names proteotypic sequences used to represent a protein or a site. Shared peptides, missed cleavage, and modification occupancy can break that mapping.
Detection scope is quadrupole isolation plus high-resolution MS/MS of product ions. Quantification uses extracted fragment ion chromatograms, not a whole-proteome intensity matrix.
Claim scope is relative contrast on the closed panel, or calibrated amount when SIS and a curve are declared. Both claims still sit on peptides unless a protein-level standard design is documented.
Peptide-level PRM quantifies scheduled surrogates. It does not by itself report molar protein concentration in tissue.
Peptide PRM Versus MRM and Versus Discovery LFQ
Route choice follows list status and claim unit.
Multiple reaction monitoring (MRM) or selected reaction monitoring (SRM) records predefined precursor-product transitions on triple quadrupole systems (Lange et al. 2008). PRM stores a high-resolution product-ion spectrum so fragments can be reviewed after acquisition (Gallien et al. 2012). Route choice after matrix pilot appears in Parallel Reaction Monitoring vs MRM: Which Fits?.
Discovery LFQ compares aligned intensities across unlabeled injections without a closed schedule. Peptide PRM fits after candidates are named. PRM principle context appears in From Principles to Applications: A Comprehensive Overview of PRM Technology.
SIS peptides calibrate the light-to-heavy ratio for the matched sequence. Spike timing must match the claim. Addition after digest does not correct unmatched extraction.
Unlabeled data-independent acquisition (DIA) or LFQ tables may supply the candidate list. Handoff requires a new targeted analysis record.
End-to-End Workflow for Peptide-Level PRM Quantification
A practical peptide PRM spine usually runs as follows.
- Freeze contrast, replicates, and relative versus amount claim.
- Close the peptide panel from discovery, literature, or orthogonal evidence.
- Select unique, LC-stable proteotypic peptides; record protein mapping rules.
- Match extraction, digest, and load across all contrast arms.
- Build retention windows and inclusion lists with realistic cycle time.
- Acquire scheduled PRM with documented isolation, collision energy, and fill-time rules.
- Extract product-ion chromatograms; review interference with multiple fragments.
- Normalize or apply SIS calibration as declared; test predefined contrast only.
- Archive panel version, method files, and software with deliverables.
Steps 1 through 4 protect the peptide surrogate. Steps 5 through 9 produce the quantitative record.
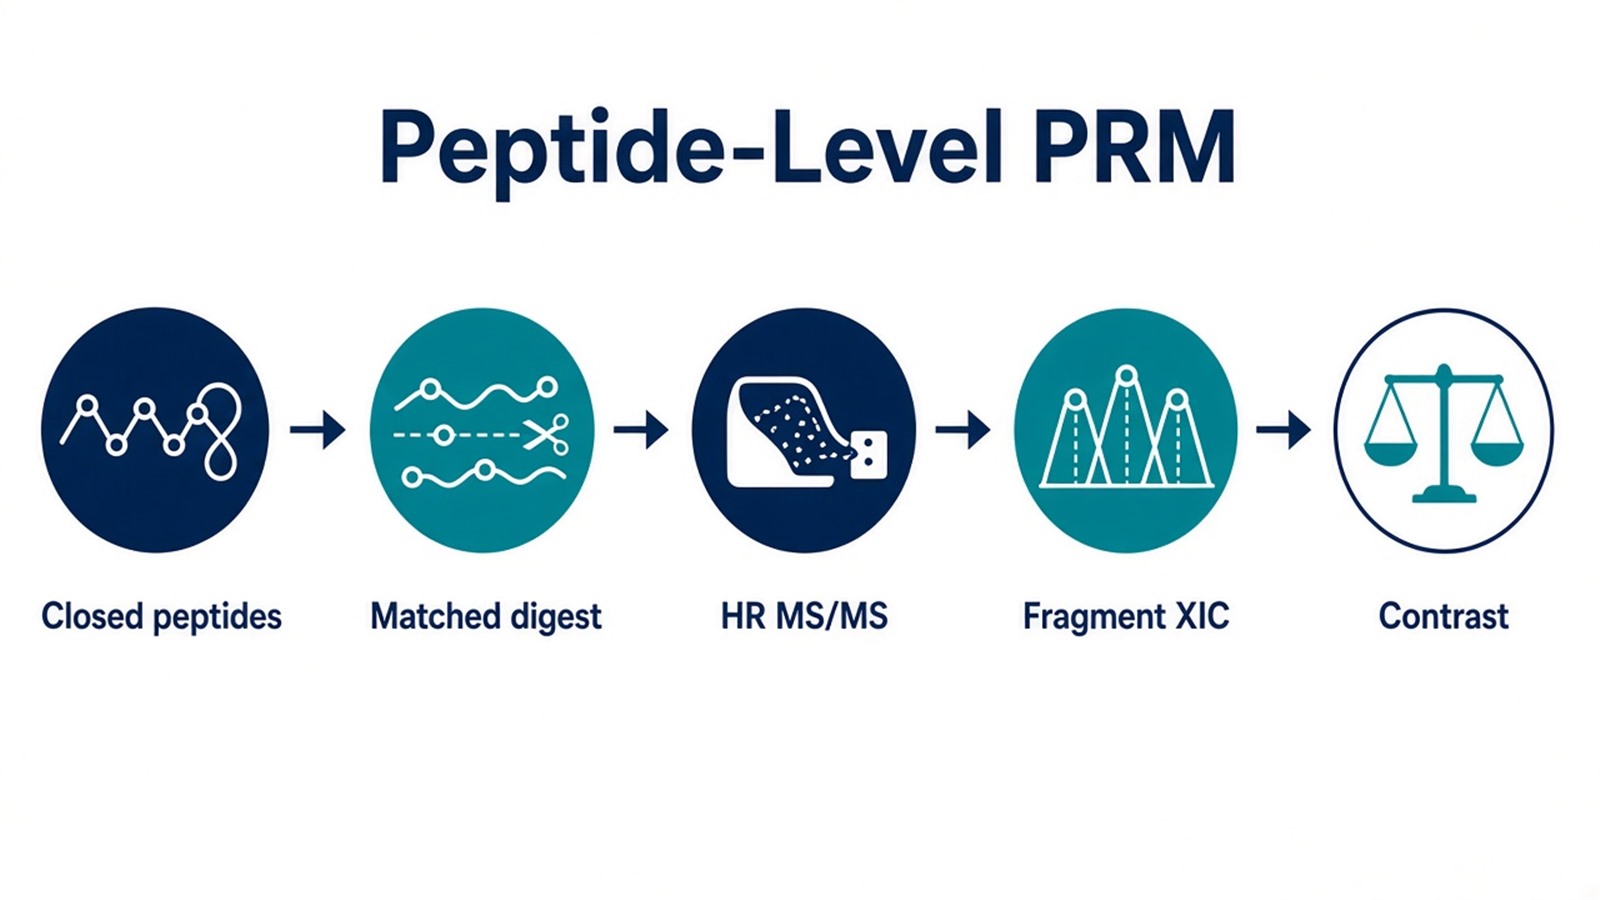
Figure 1. Peptide-level PRM depends on a closed surrogate list and matched prep before scheduled high-resolution product-ion quantification.
Parameters That Must Be Locked for Peptide PRM
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Panel |
Peptide list version and protein mapping |
Moving surrogates mid study |
Peptide rules |
Uniqueness, missed cleavage, modification policy |
Shared-peptide ambiguity |
Prep |
Matched extraction and digest |
Peptide-pool shift as biology |
Schedule |
Retention windows and cycle time |
Missing peaks at elution |
Isolation |
Precursor isolation width |
Co-isolation interference |
MS/MS |
Collision energy and resolution |
Fragment yield drift |
Quant fragments |
Extraction rules after review |
One noisy ion as the only metric |
SIS |
Spike step aligned to claim |
Peptide amount as extract amount |
Contrast |
Frozen before peak review |
Post hoc panel edits |
Software |
Extraction version archived |
Non-reproducible peaks |
Limits follow the same table. High-resolution MS/MS improves fragment review. It does not remove matrix suppression or digest variance. Panel size is bounded by peak width and cycle time. Peptide PRM does not validate drug targets or clinical biomarkers without separate study design.
How to Choose Peptide-Level PRM for Your Project
Use list status, matrix pilot, claim unit, and follow-up depth together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Closed peptide shortlist after discovery |
Peptide-level PRM |
Open LFQ as final panel |
Clean transitions already locked on triple quadrupole |
MRM or SRM |
Force PRM without interference |
Matrix interference after MRM pilot |
PRM multi-fragment review |
Whole-panel move without need |
Amount on named peptides |
PRM or MRM with SIS and curve |
Relative peaks as concentration |
Open proteome relative profiling |
Label-free LFQ or DIA |
PRM on unnamed proteins |
Intact-protein mass claim |
Not peptide PRM |
Bottom-up surrogate as intact amount |
Large panel near cycle limit |
Split panels or reduce concurrent targets |
Overfilled schedule |
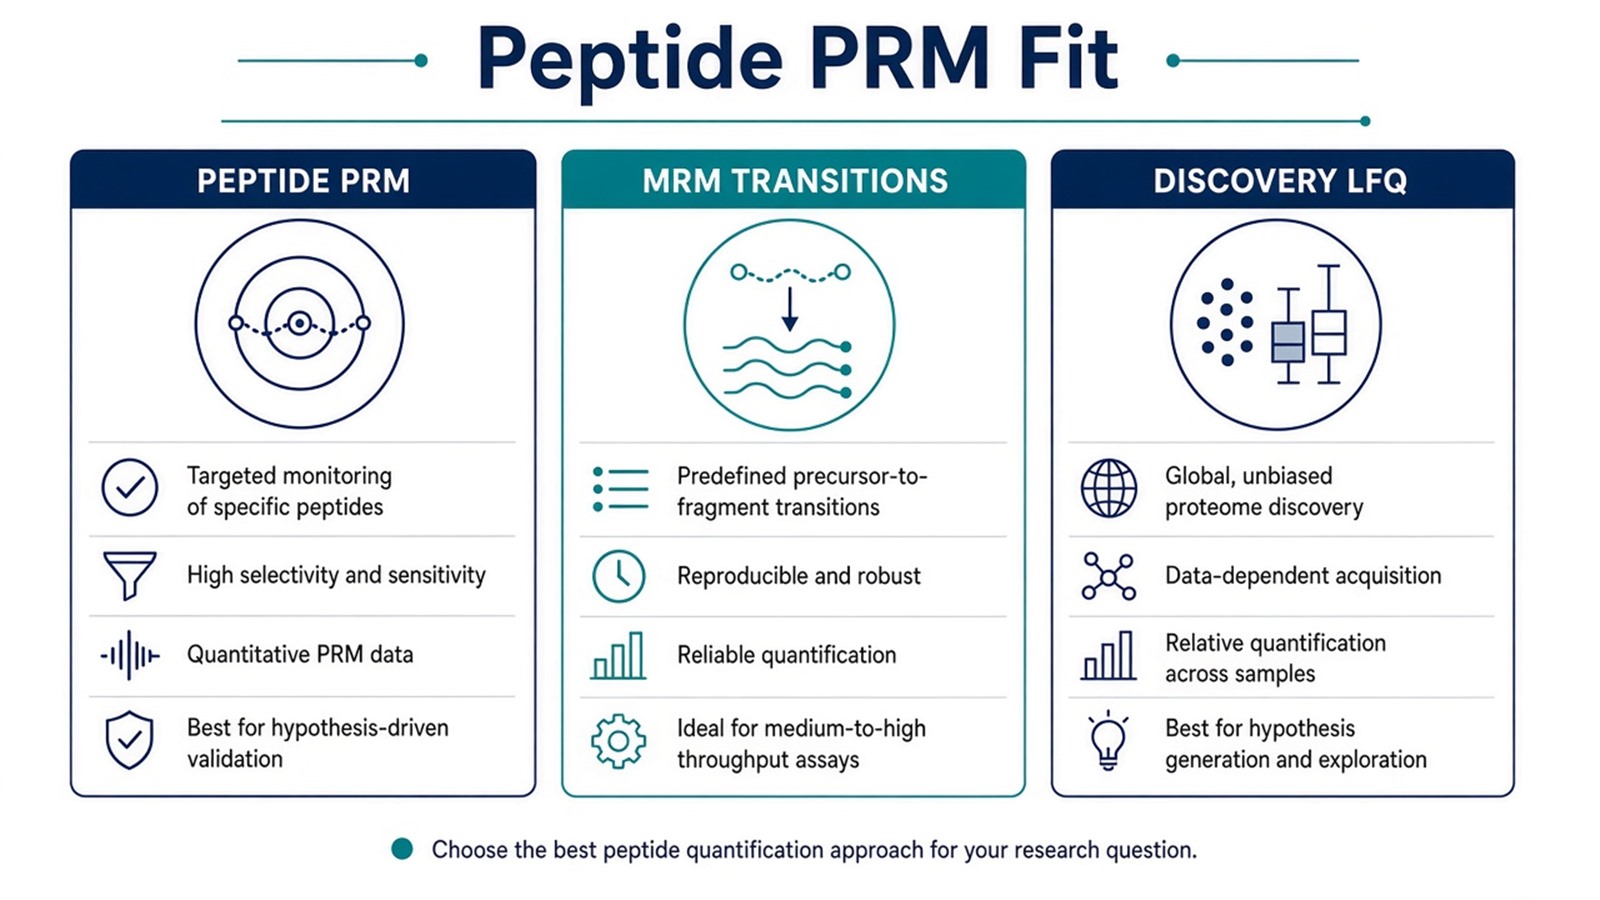
Figure 2. Choose peptide-level PRM when a closed surrogate list needs scheduled high-resolution fragment quantification; keep discovery or MRM when list status or transitions say so.
Teams with closed peptide targets, contrast map, and claim unit can review MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service when declared in the project record.
Frequently Asked Questions
1. Does peptide-level PRM equal protein concentration?
No. PRM quantifies scheduled peptides. Protein amount follows only when the surrogate mapping and standard design match that sentence.
2. Must SIS peptides be added before digestion?
Spike timing follows the claim. Addition after digest calibrates that peptide in the digest. Extraction or digestion yield of the parent protein needs a design that includes those steps.
3. Is PRM more precise than MRM for every peptide?
Not as a rule. Precision follows matrix, load, scheduling, and QC. PRM helps when multiple high-resolution fragments support assignment after MRM interference.
4. Can AI fragment picking replace a locked panel?
Learned scoring may assist review when documented. It does not replace a frozen peptide list, matched prep, or versioned methods.
5. When should projects stay on discovery instead of peptide PRM?
When proteins are not yet named or the brief needs proteome-wide relative profiling, LFQ or DIA remains the matching block.
MtoZ Biolabs can discuss peptide-level PRM when panel version, matrix prep, and contrast design are documented.
If further evaluation of PRM versus MRM, SIS calibration need, or discovery-to-PRM handoff is required, project details can be submitted for a feasibility review.
Reference
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S. Gallien, E. Duriez, A. Crone, M. Kellmann, T. Moehring, T. Domon (2012). Targeted proteomic quantification on quadrupole-orbitrap mass spectrometers. Mol. Cell. Proteomics, 11, 1709-1723. https://doi.org/10.1074/mcp.O112.019232
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
