



Affinity purification-mass spectrometry (AP-MS) asks which proteins remain with a tagged bait after affinity-handle capture, washing, and LC-MS/MS identification from a defined expression system. Immunoprecipitation-mass spectrometry (IP-MS) asks the same enrichment question for an endogenous bait recovered with a target-specific antibody. Both workflows return candidate co-complex members, not proof of binary direct binding (Hein et al. 2015). Prefer IP-MS when native expression, tissue or primary-cell context, and an IP-competent antibody are available. Prefer AP-MS when tagging is acceptable, antibody performance is weak or unknown, or matched construct series must share one purification chemistry. Detection stops at enrichment versus handle-matched controls after bait recovery is confirmed.
Researchers who already have a named bait, antibody or tag status, and a written interactome claim can also review the MtoZ Biolabs Protein-Protein Interaction Analysis and MS-Based Protein-Protein Interaction Analysis Service for project-specific workflow feasibility.
What Each Interactome Workflow Is Built to Answer
IP-MS and AP-MS are often treated as interchangeable names for the same interactome experiment. They are not. Both enrich a bait and identify co-purifying proteins by LC-MS/MS. They start from different biological materials and fail for different reasons.
IP-MS captures an endogenous bait with a specific antibody. Its success depends on whether that antibody works under the planned lysis and wash conditions. Western blot reactivity alone is not enough. The antibody should show bait enrichment, acceptable background under IP conditions, and recovery compatible with LC-MS/MS.
AP-MS, in the usual interactome-workflow sense, captures a tagged bait through an affinity handle such as FLAG, HA, His, Strep, or GFP-related systems. Its success depends on tag design, expression level, and empty-tag controls rather than on a target-specific IP antibody. Primary cells, tissues, and many clinical specimens cannot accept a tag, so those matrices usually stay on IP-MS when an antibody exists.
Neither method proves a direct physical contact. Both return candidate interactors that require quantitative filtering and orthogonal validation. Immobilized recombinant bait capture is a different assay and is compared in Endogenous IP-MS vs. Tagged Pull-Down MS.
Antibody-based intake is scoped through the IP-Mass Spectrometry Service. Fusion or immobilized bait follow-up is scoped through the Fusion Protein Interaction Analysis Service | Pull-Down and MS.
IP-MS and AP-MS differ in how the bait enters the experiment, not in the mass spectrometer used for identification.
Bait Enrichment, Background Proteins, and Control Design
High-confidence interactome lists fail more often at bait recovery than at the database search. If bait peptides are weak or missing, partner names in the same file are not interpretable as a neighborhood. Relative intensity values can compare the same protein across matched enrichments. They do not convert a name into a tighter binder or a binary contact.
Background proteins are expected. IP-MS background is dominated by antibody constant-region binders, bead binders, and off-target antibody capture. AP-MS background is dominated by tag-binding proteins, resin binders, and partners recruited by overexpression or non-native localization. A contaminant frequency catalog such as the CRAPome is a prior on how often a protein appears in control purifications, not a verdict that a name is false in the current matrix (Mellacheruvu et al. 2013).
Controls must match the capture handle.
|
Control class |
IP-MS |
AP-MS |
What it can support |
|---|---|---|---|
|
Empty bead or resin |
Bead binders |
Bead binders |
Subtract resin-only proteins |
|
Handle control |
Isotype IgG |
Empty-tag line |
Subtract antibody or tag binders |
|
Bait dependence |
Knockdown or knockout when feasible |
Untagged parental line |
Test bait-specific enrichment |
|
Expression match |
Native abundance |
Matched tag expression |
Reduce overexpression bias |
Wash stringency is part of the method. Harsh washes can drop weak or transient partners. Mild washes retain more candidates and more background. The wash recipe belongs next to the partner table.
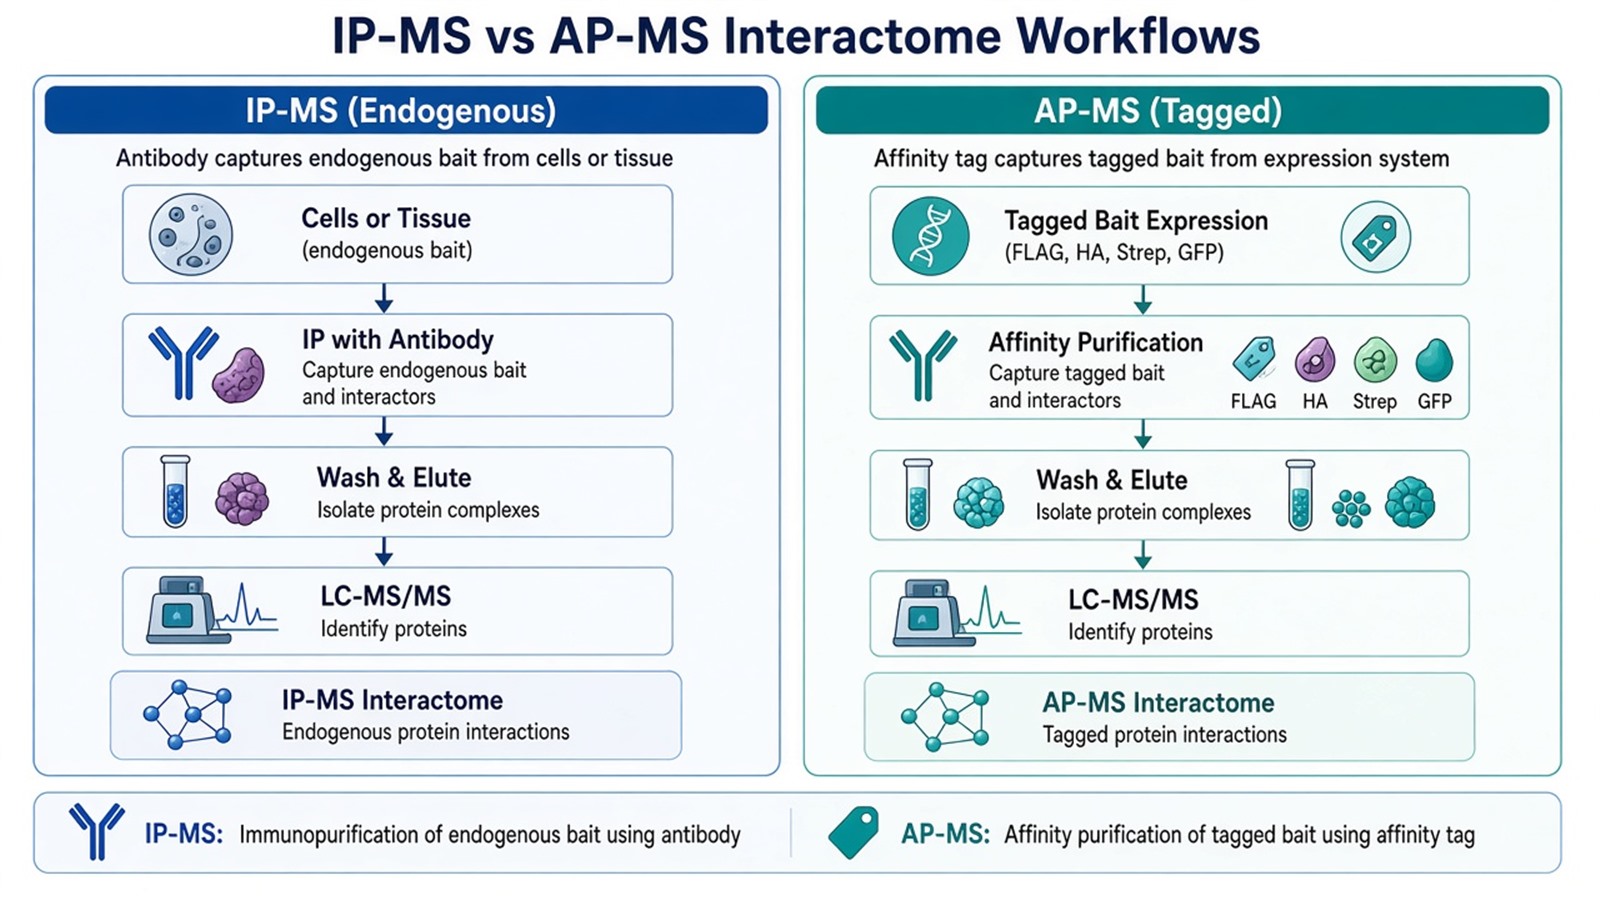
Figure 1. IP-MS depends on antibody capture of an endogenous bait; AP-MS depends on affinity-tag purification of a designed construct.
Filtering High-Confidence Interactors
Identification of a protein in an IP-MS or AP-MS eluate is not a confidence call. Ranking requires bait recovery, handle-matched controls, and a declared enrichment rule. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). Those scores still operate on enrichment tables. They do not convert a co-purified name into a direct binder or a cellular occupancy.
A practical filtering sequence is:
- Confirm bait peptides and expected bait coverage in the eluate.
- Remove proteins that fail identification or quantitation QC.
- Rank remaining names against handle-matched controls, not against an unmatched public list alone.
- Flag high-frequency bead, antibody, or tag contaminants for extra scrutiny.
- Keep a shortlist for reciprocal capture, defined-pair pull-down, or another orthogonal test.
Single unreplicated enrichments are weak evidence. Replicates and a predefined candidate filter should be set before acquisition whenever possible. Pathway maps interpret a shortlist. They do not repair a missing empty-tag line or a failed antibody IP.
For how enrichment lists constrain discovery without predicting structure, see How Can Mass Spectrometry Be Used to Analyze or Predict Protein-Protein Interactions?. Screening-level method choice is covered in Measuring Protein-Protein Interactions.
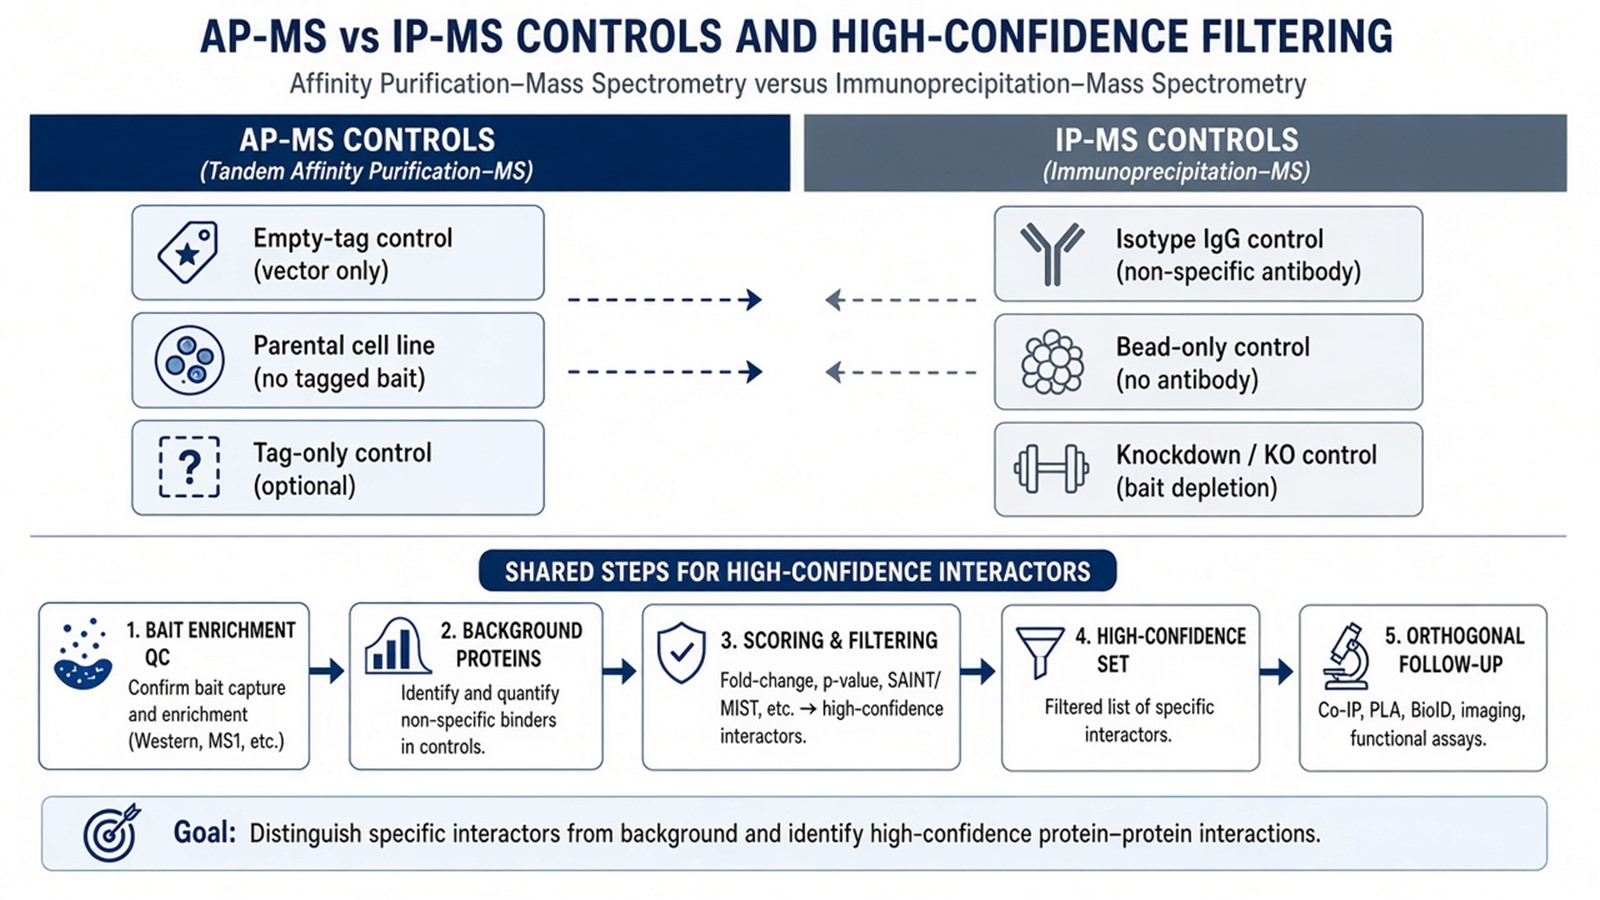
Figure 2. IP-MS and AP-MS need different handle controls; both still require bait recovery before a partner is treated as high confidence.
Judgment Criteria: Choosing IP-MS or AP-MS
|
Decision factor |
IP-MS |
AP-MS |
|---|---|---|
|
Bait format |
Endogenous protein |
Tagged protein in an expression system |
|
Capture reagent |
Target-specific antibody |
Affinity resin for the tag |
|
Needs IP-competent antibody |
Yes |
No |
|
Endogenous context |
Strong when the antibody works |
Depends on expression and tag design |
|
Best for |
Native bait interactomes in cells or tissue |
Systematic tagged interactomes and construct series |
|
Main risk |
Poor antibody performance and antibody background |
Tag effects and overexpression artifacts |
|
Typical next step |
Reciprocal IP or functional tests |
Filter tag background, then endogenous confirmation |
Prefer IP-MS when the bait must remain endogenous and an antibody with documented IP performance in the planned matrix is available. Typical fits include tissues, primary cells, or specimens that cannot be tagged, studies in which overexpression would distort partner selection, and projects that need the bait in its native modification state.
Prefer AP-MS when tagging is acceptable and antibody performance is the bottleneck. Typical fits include mutant, truncation, or domain-swap comparisons under one purification chemistry, condition series that require matched bait recovery, and targets with no antibody validated for IP.
Reassess IP-MS when available antibodies fail under wash conditions, cross-react with abundant off-targets, or cannot provide reproducible bait recovery. Reassess AP-MS when endogenous stoichiometry is essential, when tagging is prohibited by the sample type, or when the tag changes localization or function.
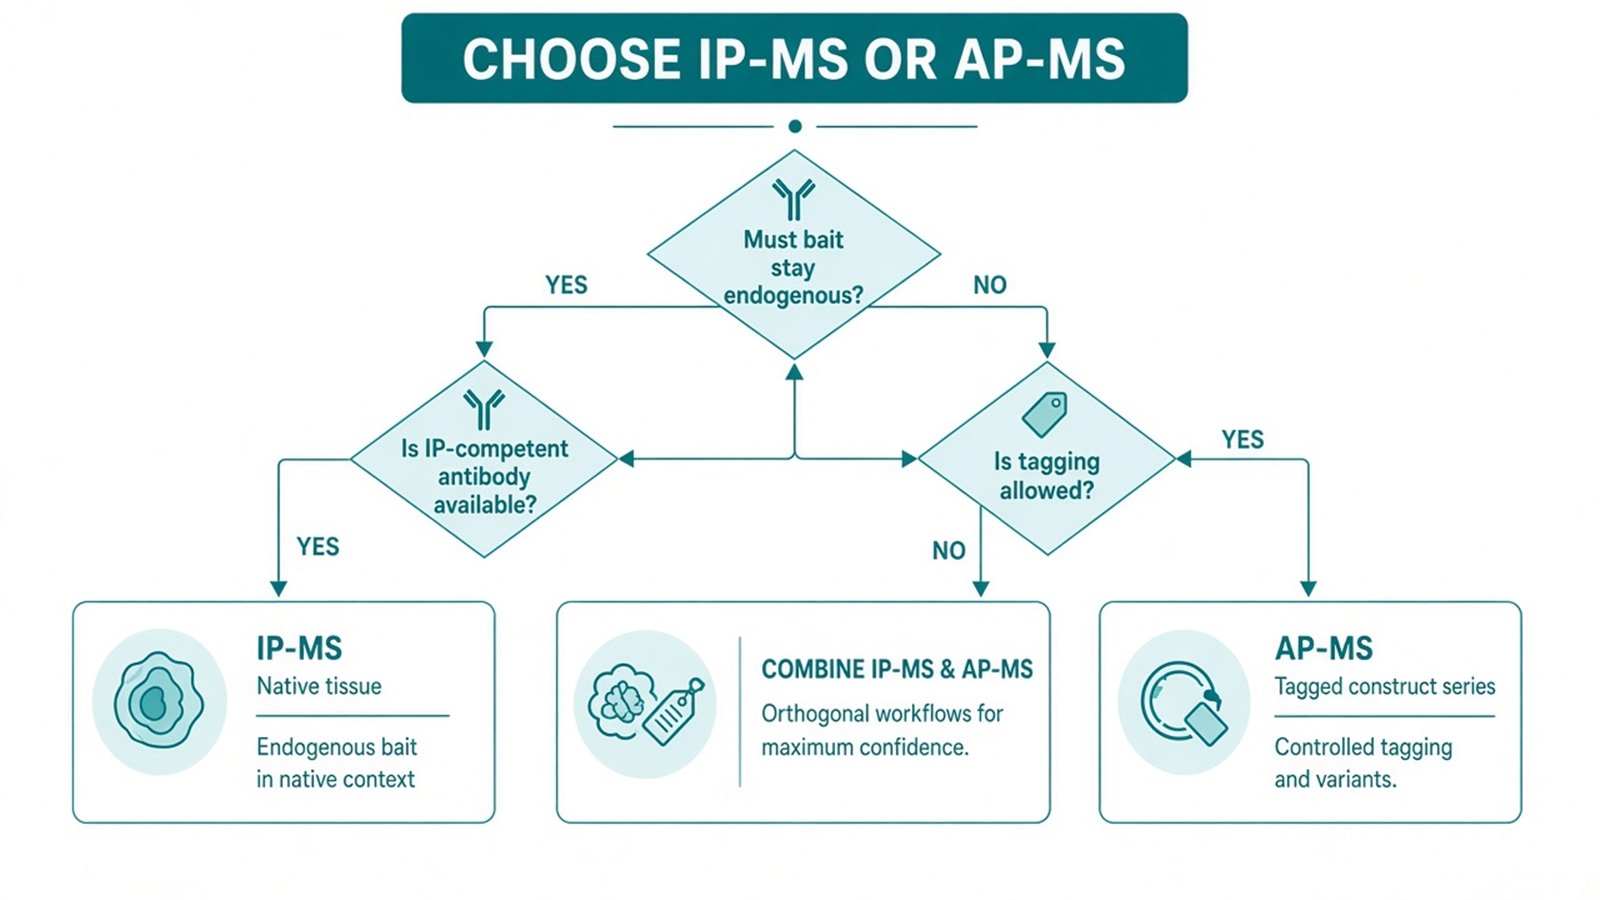
Figure 3. Antibody readiness and tagging acceptability are the primary branch points between IP-MS and AP-MS.
Common Biases and How to Read the Result
IP-MS tends to overcall proteins that bind antibody constant regions, proteins that stick to beads under the chosen lysis chemistry, and proteins related to antibody cross-reactivity. Weak bait recovery can also create noisy lists dominated by contaminants.
AP-MS tends to overcall proteins that bind the affinity tag or resin, partners recruited by overexpression, and interactions stabilized by non-native localization of the tagged bait. Highly expressed tagged baits can recover sticky or abundant proteins that are not meaningful partners.
After scoring, a protein-level value can support a relative statement: protein B is enriched with bait A versus the matched control inside this capture recipe. Peptide support, bait recovery, control frequency, and wash conditions belong next to that statement. Ordinary interactome tables should not rank different proteins as tighter binders by intensity alone, infer a diagnostic interaction, or transfer a partner list from one lysis buffer to another study as a conserved complex.
An IP-MS or AP-MS table supports enrichment of named proteins with a captured bait under stated controls, not a certified binary interactome.
When Both Workflows Belong in One Program
Some programs discover candidates by AP-MS, then test endogenous recovery by IP-MS when a suitable antibody becomes available. Others qualify an antibody by IP-MS and later use AP-MS for mutant or condition series under matched tag chemistry. Overlapping but non-identical shortlists are expected because expression context, wash chemistry, and capture handles differ.
Pull-down-MS is not a third name for AP-MS. It typically uses an immobilized recombinant bait to test retention under controlled conditions. Cellular AP-MS recovers proteins associated with a tagged bait from an expression system. Choose the assay that matches the claim, then sequence confirmation rather than forcing one enrichment to answer every question.
Once bait format, antibody or tag readiness, and control set are defined, the project can be scoped as IP-MS, AP-MS, or a staged interactome sequence. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service and GST Pull Down Assay Service for sample evaluation and capture planning.
Frequently Asked Questions
1. Is AP-MS always better when no good antibody exists?
AP-MS is often the more productive discovery route when tagging is acceptable. If tagging is not allowed, antibody improvement, recombinant pull-down, or a non-MS interaction assay may be required instead.
2. Does IP-MS require an antibody already shown to work in IP?
For interpretable discovery work, yes in practical terms. An antibody that only works on a Western blot can produce a weak IP-MS dataset with poor bait recovery.
3. Can AP-MS and IP-MS be used in the same project?
Yes. Tagged discovery and endogenous confirmation answer different claims. They are complementary when both the handle and the antibody can be controlled.
4. How is pull-down-MS different from AP-MS?
Pull-down-MS typically uses an immobilized recombinant bait to test binding under controlled conditions. AP-MS usually purifies a tagged bait from a cellular expression system and recovers proteins associated in that context.
5. Which method is better for disease or tissue samples?
IP-MS is usually closer to the biology when endogenous bait from tissue or primary material must be studied and an IP-competent antibody is available. AP-MS is stronger when engineered models can carry the tagged bait.
Reference
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
